
21 CFR Part 820 as amended – the Quality Management System Regulation – has been in force since 2 February 2026. It incorporates ISO 13485:2016 by reference and adds a small, specific FDA layer on top. This QMSR documentation set documents that layer, and leaves the quality system itself to ISO 13485.
On this page:
- Why an ISO 13485 certificate is not enough
- What is inside the QMSR documentation pack
- Built on the inspection programme FDA actually uses
- Who the QMSR documentation pack is written for
- What the QMSR documentation pack does not do
- Frequently asked questions
- Related toolkits
Why an ISO 13485 certificate is not enough
A certificate evidences conformity with the standard. It does not evidence the four bridges named in 820.10(b) from ISO clauses to Parts 830, 821, 803 and 806; the design-control applicability determinations in 820.10(c) and (d); the seven complaint data elements and six servicing data elements in 820.35; the recorded UDI required by 820.35(c) as opposed to the printed one; the five accuracy checks and documented release in 820.45; or the FDA definitions in 820.3 that supersede the ISO wording – including Organization, which here means manufacturer. Those are the gaps this pack closes.
There is one change most quality systems have not caught up with. Under the former Quality System Regulation, section 820.180(c) exempted management review reports, internal quality audit reports and supplier audit reports from FDA review; an investigator could require only a written certification that they had been performed. The QMSR has no section 820.180. The exemption went with it, and Compliance Program 7382.850 – which replaced QSIT on the same day – lists Internal Audits and Management Review among the elements an investigator may select. Those reports can now be read.
Two documents in this QMSR documentation set exist solely for that change, and it is the single most common blind spot in a QSR-era quality system.
What is inside the QMSR documentation pack
73 documents – 45 Word and 28 Excel – in nine sections that follow the transition rather than the old regulation order.
- Programme foundation – 6 documents establishing scope, definitions and the FDA layer itself.
- QSR to QMSR transition – 7 documents, including all 31 sections of the old regulation mapped to where each requirement now lives.
- Quality management system requirement – 8 documents covering 820.10 and the four bridges.
- Design and development applicability – 6 documents for the 820.10(c) and (d) determinations.
- Control of records – 9 documents, including the confidentiality and FDA-review position after the loss of 820.180.
- Labeling and packaging controls – 8 documents covering 820.45 and its five accuracy checks.
- Cross-referenced FDA parts – 10 documents for Parts 830, 821, 803 and 806.
- Inspection readiness and evidence – 12 documents built on the inspection programme FDA actually uses.
- Assessment and mapping – 7 documents, including the master cross-reference matrix.
Built on the inspection programme FDA actually uses
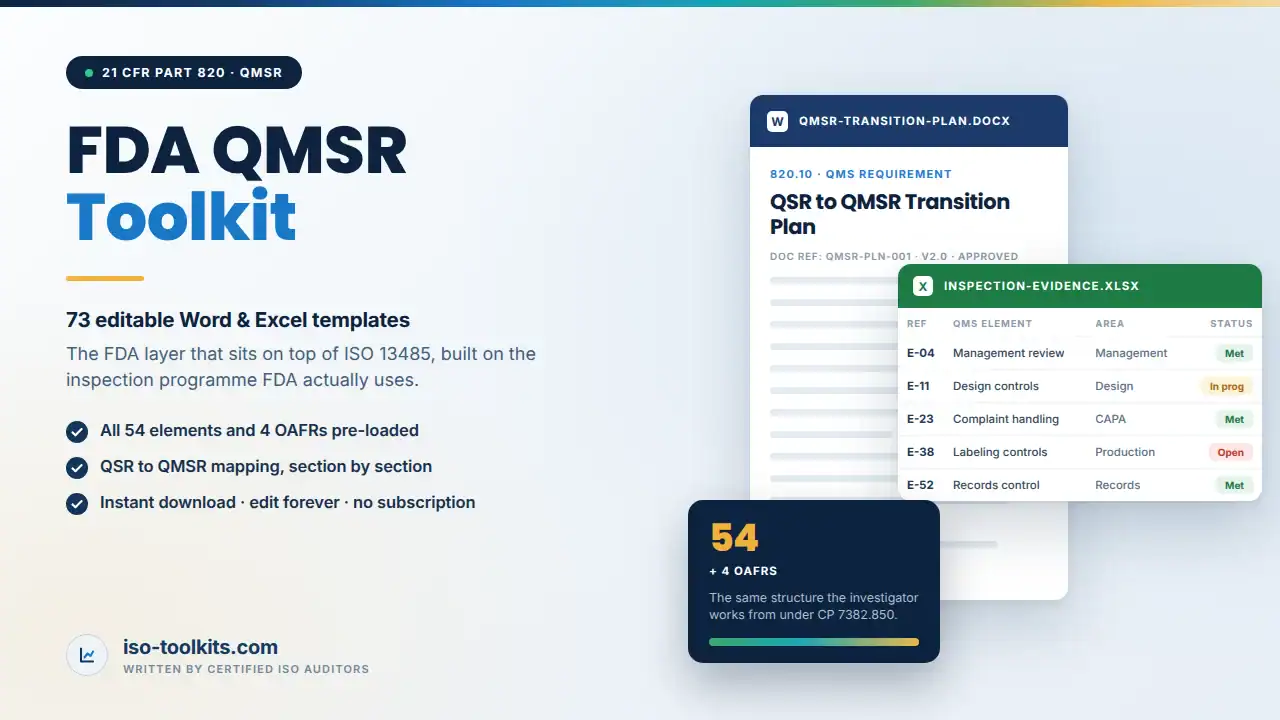
Since 2 February 2026, FDA inspects under Compliance Program 7382.850, not QSIT. It organises the quality system into 54 elements across six QMS Areas, plus 4 Other Applicable FDA Requirements, each mapped to ISO 13485 clauses and 820 sections.
Four workbooks come pre-loaded with all 54 elements and all 4 OAFRs – the evidence map, the audit checklist, the conformance register and the master cross-reference matrix. You prepare against the same structure the investigator is working from, which is a materially different exercise from preparing against a generic quality manual.
Twelve of the 28 workbooks ship seeded rather than empty. The seven complaint data elements and six servicing data elements appear as actual columns, so a missing field is visible rather than buried in narrative. The five class I device types named in 820.10(c)(2) are listed so you can check your portfolio against them directly. That is the difference between QMSR documentation you fill in and a template you have to design first.
Who the QMSR documentation pack is written for
- Device manufacturers marketing in the United States, at any class.
- Specification developers, contract manufacturers, relabelers, repackers and remanufacturers – all manufacturers under 820.3(b).
- Quality and regulatory teams transitioning a QSR-era quality system.
- Organisations preparing for a first inspection under Compliance Program 7382.850.
What the QMSR documentation pack does not do
It does not make you compliant and it does not replace an ISO 13485 quality management system. If you need the quality system itself, that is the ISO 13485 pack and the two are built to sit together.
It is not a submission pack. Premarket work is a separate exercise.
It does not cover EU MDR or IVDR. Those are separate regimes with their own technical documentation.
Frequently asked questions
We are already ISO 13485 certified. Do we need this?
Yes, if you market in the US. The certificate does not evidence the FDA-specific layer – the 820.10(b) bridges, the 820.35 data elements, the recorded UDI, the 820.45 checks or the 820.3 definitions that supersede ISO wording.
What changed about audit and management review reports?
The QMSR has no section 820.180, so the exemption that kept those reports out of FDA review is gone. Compliance Program 7382.850 lists both among the elements an investigator may select.
Does this replace our quality manual?
No. It documents the FDA layer on top of it. Buy the ISO 13485 pack as well if you do not have a quality system.
What formats are the files in?
Native Microsoft Word and Excel, fully editable, with placeholders marked throughout.
Related toolkits
It is built to sit on top of the ISO 13485 toolkit, with the ISO 14971 toolkit for the risk management file. Manufacturers also selling into the EU need the EU MDR toolkit. FDA publishes the regulation and its FAQ at fda.gov.
Implementing for clients? The Consultant Package licenses all 86 toolkits and assessment tools on this site for unlimited client engagements, under one firm-wide licence. One payment of $1,399, no subscription and no per-client fee.
Delivery, format and licence
The QMSR documentation pack downloads immediately after checkout as native Microsoft Word and Excel files. Nothing is locked, nothing is a PDF you cannot edit, and no add-on or portal login is needed to open it. Every organisation-specific value is marked as a placeholder so you can see what still has to be decided.
One payment, no subscription and no annual renewal. The source files behind the QMSR documentation pack are yours to adapt for your own organisation for as long as you need them, including future revisions of your own documents.


Reviews
There are no reviews yet.