
The Medical Device Regulation is not a standard you implement once. It has been amended eight times, consolidated eight times, and is served by 125 MDCG guidance documents and a harmonised standards list amended roughly twice a year. That is why most MDR technical documentation is quietly out of date.
On this page:
- EUDAMED is not a future project, and three other dates that have already passed
- What is inside the MDR technical documentation pack
- Written once, or written per device, and the difference is marked
- Who the MDR technical documentation pack is written for
- What the MDR technical documentation pack does not do
- Frequently asked questions
- Related toolkits
EUDAMED is not a future project, and three other dates that have already passed
Under Commission Decision (EU) 2025/2371 and the six-month clock in Article 123(3)(d), four EUDAMED modules became mandatory on 28 May 2026: actor registration, UDI and device registration, notified bodies and certificates, and market surveillance. Three of those are direct manufacturer obligations and they are live now. A manufacturer without a Single Registration Number is not preparing for something – Article 31(7) makes registration data publicly accessible, so the gap is visible to customers, competitors and competent authorities at the same time.
Two Commission Delegated Regulations of 20 March 2026 took effect in the July consolidation. 2026/1359 sets the list of class IIb implantable devices exempted from per-device technical documentation assessment under Article 52(5); 2026/1451 amends the Article 61(6)(b) list of devices exempted from clinical investigations. Both change what manufacturers actually have to do, and both are reflected here in one dated supplement rather than frozen into forty documents.
The Article 120 transition conditions are the third trap. The extension to 2027 or 2028 was conditional on things that had to happen in 2024 – a quality management system meeting Article 10(9) and a formal notified body application by 26 May 2024, and a signed written agreement by 26 September 2024. If a condition was missed, the extension never applied to that device. The transition tracker in this MDR technical documentation takes evidence references rather than a Yes against each of the five conditions.
What is inside the MDR technical documentation pack
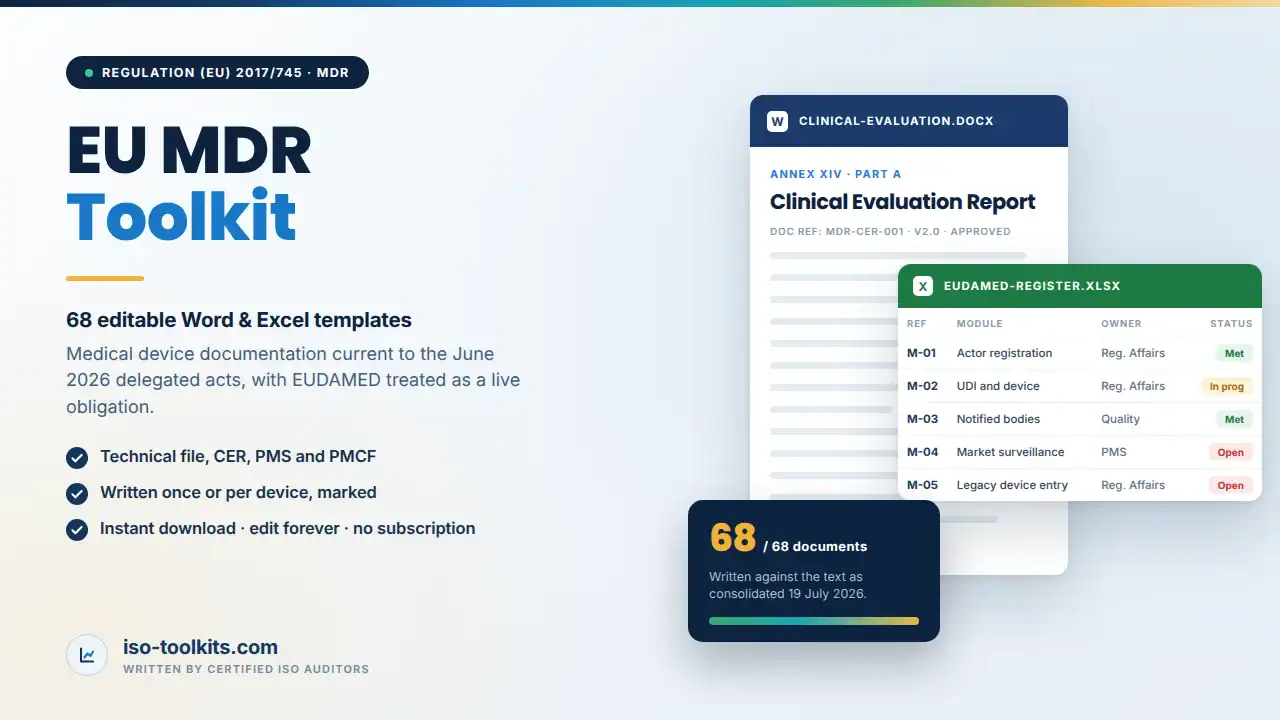
68 editable templates across the full regulatory scope, each stating the consolidation date of the text it was written against.
- Technical documentation – the Annex II and Annex III structure, written once or per device as the Regulation requires, with each document marked accordingly.
- Clinical evaluation – the CER structure, the clinical evaluation plan, the literature method and the equivalence justification.
- PMS and PMCF – the post-market surveillance plan, the PMCF plan and evaluation report, the PSUR and the trend reporting procedure.
- Vigilance – incident and field safety corrective action reporting, on the MDR definitions rather than the old directive ones.
- UDI and registration – UDI assignment, the EUDAMED actor and device registration procedures, and the SRN record.
- Conformity assessment – route selection reflecting the March 2026 delegated acts, and the declaration of conformity.
- Economic operators – authorised representative, importer and distributor obligations, and the PRRC appointment.
- Transition – the Article 120 tracker on evidence references, including Article 120(3d) which applies PMS, vigilance and registration to legacy devices today.
- Harmonised standards – the position stated correctly, including which standards are and are not harmonised under the MDR.
Written once, or written per device, and the difference is marked
The structural error that costs the most is a single shared file used as evidence for a portfolio. The Regulation requires some things of the organisation and some things of each device, and a technical file that merges the two cannot serve as evidence for any individual device.
Every document in this MDR technical documentation set is marked as written once, written per device, or both. That single distinction decides how a file is assembled and is the first thing a notified body reviewer notices when it has been ignored.
The same care applies to what is not harmonised. IEC 62304, IEC 62366-1 and EN ISO 20417 are widely described as harmonised under the MDR. The pack states the actual position, because a technical file that claims presumption of conformity it does not have is a finding waiting to happen.
Who the MDR technical documentation pack is written for
- Manufacturers placing medical devices on the EU market, at any class.
- Authorised representatives, importers and distributors with their own MDR obligations.
- Quality and regulatory teams whose documentation predates the May 2026 EUDAMED modules.
- Organisations running legacy devices under Article 120 that need the conditions evidenced rather than assumed.
What the MDR technical documentation pack does not do
It is not a quality management system. MDR expects ISO 13485 underneath it, and the two packs are built to sit together.
It does not write your clinical evidence. It gives the structure, the method and the plan; the data is yours.
It is not the IVD pack. IVDR article numbers differ and the two regulations diverge exactly where it costs most – classification, clinical evidence and the technical file structure.
Frequently asked questions
Is this the same as the IVDR pack?
No. If you make in vitro diagnostics you want the IVDR pack; the article numbers and the evidence requirements are different.
Do we still need ISO 13485?
In practice yes. MDR requires a quality management system and ISO 13485 is how almost everyone demonstrates it. Buy them together if you do not already have one.
Is the EUDAMED position current?
Yes – it is written against the four modules that became mandatory on 28 May 2026, including the staged follow-on obligations.
What formats are the files in?
Native Microsoft Word and Excel, fully editable, with placeholders marked throughout.
Related toolkits
It is normally bought with the ISO 13485 toolkit for the quality system and the ISO 14971 toolkit for the risk management file Annex I requires. Manufacturers of in vitro diagnostics need the EU IVDR toolkit instead. The consolidated Regulation is on EUR-Lex.
Implementing for clients? The Consultant Package licenses all 86 toolkits and assessment tools on this site for unlimited client engagements, under one firm-wide licence. One payment of $1,399, no subscription and no per-client fee.
Delivery, format and licence
The MDR technical documentation pack downloads immediately after checkout as native Microsoft Word and Excel files. Nothing is locked, nothing is a PDF you cannot edit, and no add-on or portal login is needed to open it. Every organisation-specific value is marked as a placeholder so you can see what still has to be decided.
One payment, no subscription and no annual renewal. The source files behind the MDR technical documentation pack are yours to adapt for your own organisation for as long as you need them, including future revisions of your own documents.


Reviews
There are no reviews yet.